Research themes
My primary research interests currently focus on methodological developments in ab initio Quantum Monte Carlo (QMC) methods and implementing these developed theories into versatile computational software. The quantum mechanical behavior of electrons, which governs the chemical and physical properties of materials, can, in principle, be predicted by solving the Schrödinger equation. However, except for special cases, analytical solutions are unattainable. Therefore, numerical solutions obtained via ab initio calculations using high-performance computing resources are typically employed.
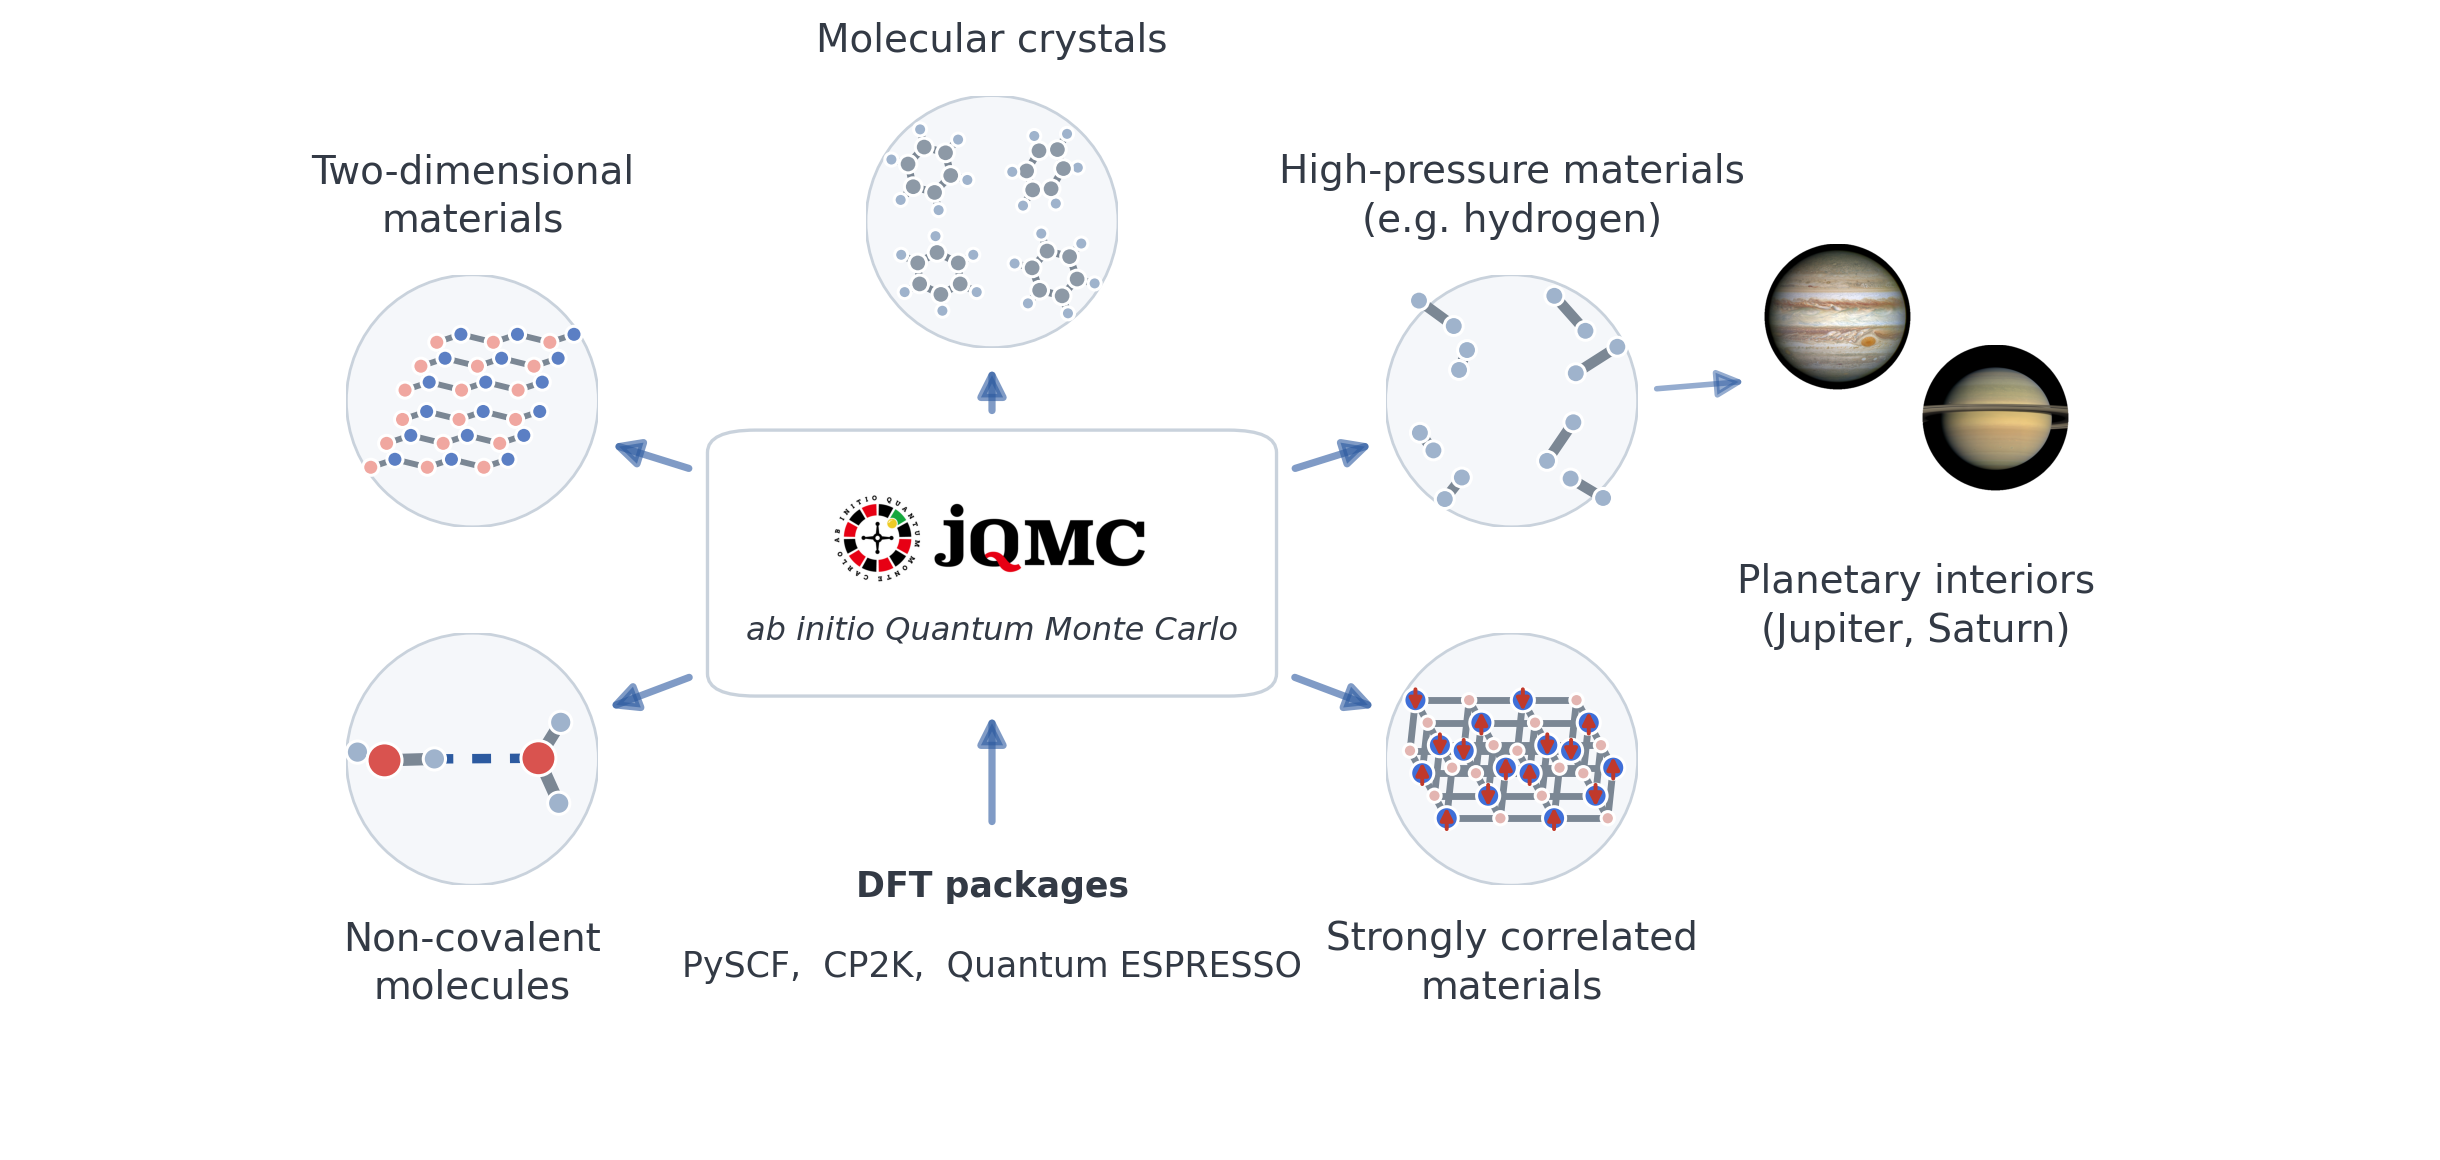
What we compute: the systems our ab initio QMC codes, jQMC and TurboRVB, are applied to - two-dimensional materials, molecular crystals, non-covalent molecules, semiconductors and insulators, strongly correlated materials, and high-pressure materials such as dense hydrogen, whose equation of state in turn determines the interior structure of the giant planets. (Planets: Jupiter by NASA, ESA and A. Simon (GSFC), Saturn by NASA / JPL-Caltech / Space Science Institute, both public domain.)
At present, wavefunction theory and density functional theory (DFT) represent the most widely used first-principles computational approaches, achieving notable successes across quantum chemistry, materials science, and condensed matter physics. Nevertheless, wavefunction-based methods suffer from difficulties in handling periodic boundary conditions and poor computational scaling with electron numbers. Meanwhile, DFT faces significant issues arising from the arbitrary choice of exchange-correlation functionals. Thus, there is a strong demand for developing more rigorous, next-generation electronic structure calculation methods.
Ab initio Quantum Monte Carlo (QMC) methods numerically solve the many-body Schrödinger equation using Monte Carlo integration techniques. Among current electronic structure methods, QMC provides solutions that are closest to the exact solution. Unlike DFT, QMC eliminates the arbitrariness associated with selecting exchange-correlation functionals and readily facilitates parallel computing, maximizing the potential of modern high-performance computing resources. Consequently, QMC possesses disruptive innovation potential and is capable of replacing existing electronic structure calculation methods.
Historically, the computational cost of QMC methods has limited their practical applicability to realistic material systems of interest. However, the recent availability of exascale-class supercomputers, such as the Fugaku supercomputer at Japan’s RIKEN institute, has made it feasible to conduct practical QMC calculations within realistic computation times. Nevertheless, due to methodological limitations and lacking well-established general-purpose software, QMC has not yet supplanted conventional wavefunction and DFT methods. Many research groups are actively working on novel methodologies and corresponding software implementations to address these issues.
Currently, I am developing two ab initio QMC packages central to my research activities:
jQMC (our new flagship code), [link to GitHub], which is an ab initio quantum Monte Carlo (QMC) simulation package developed entirely from scratch using Python and JAX. Originally designed for molecular systems –with future extensions planned for periodic systems– jQMC implements two well-established QMC algorithms: Variational Monte Carlo (VMC) and a robust and efficient variant of Diffusion Monte Carlo algorithm known as Lattice Regularized Diffusion Monte Carlo (LRDMC). By leveraging JAX just-in-time (jit) compilation and vectorized mapping (vmap) functionalities, jQMC achieves high-performance computations especially on GPUs while remaining portable across CPUs, GPUs, and TPUs. See here for the details of JAX.
TurboRVB (the established Fortran90 code), [link to GitHub], which implements the core computational kernels for ab initio QMC calculations. Originally initiatedabout two decades ago by Professor Sandro Sorella (International School for Advanced Studies, Italy) and Prof. Michele Casula (CNRS, Sorbonne University, France), TurboRVB has benefited from the contributions of numerous researchers. I joined its development team in 2017 and currently serve as its primary developer and maintainer following Professor Sorella’s passing. TurboRVB, written in Fortran90, is undergoing refactoring to adopt modern coding practices (new contributors are welcome!).
Below are a few examples of our recent methodological developments:
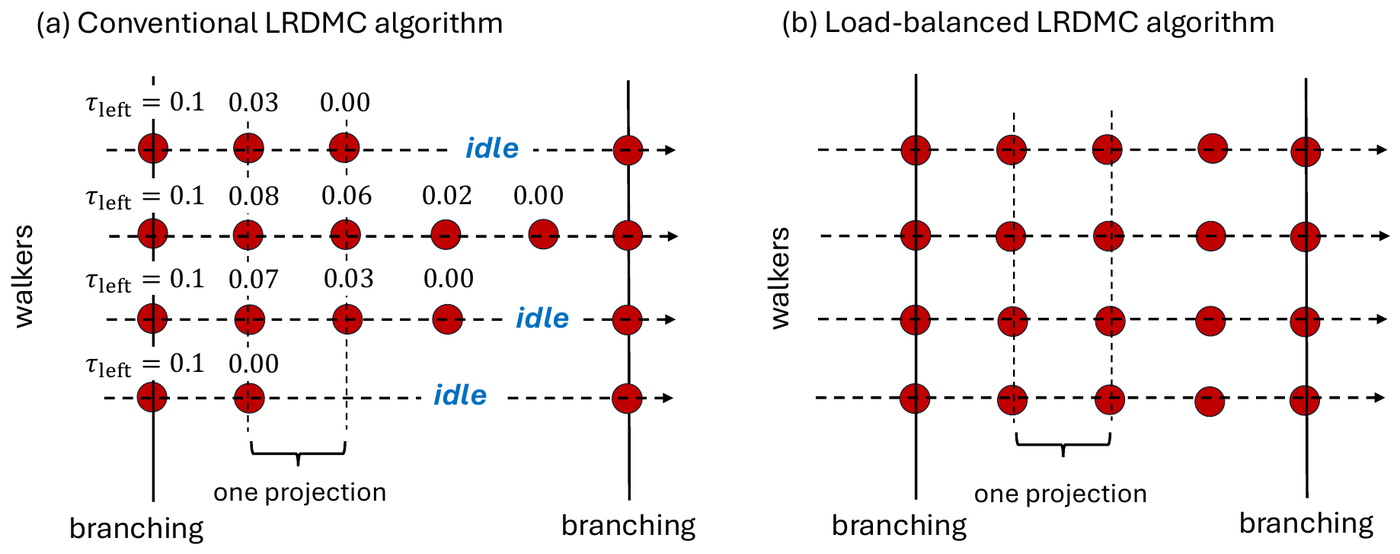
Load balancing in the lattice regularized diffusion Monte Carlo (LRDMC) algorithm. In the conventional algorithm (a), each walker performs a different number of projection steps between two branching events, which leaves many walkers idle on a parallel machine. In our load-balanced algorithm (b), every walker performs the same number of projections, so the parallel efficiency is preserved. Reproduced from K. Nakano, S. Sorella, and M. Casula, J. Chem. Phys. 163, 194117 (2025), licensed under CC BY 4.0.
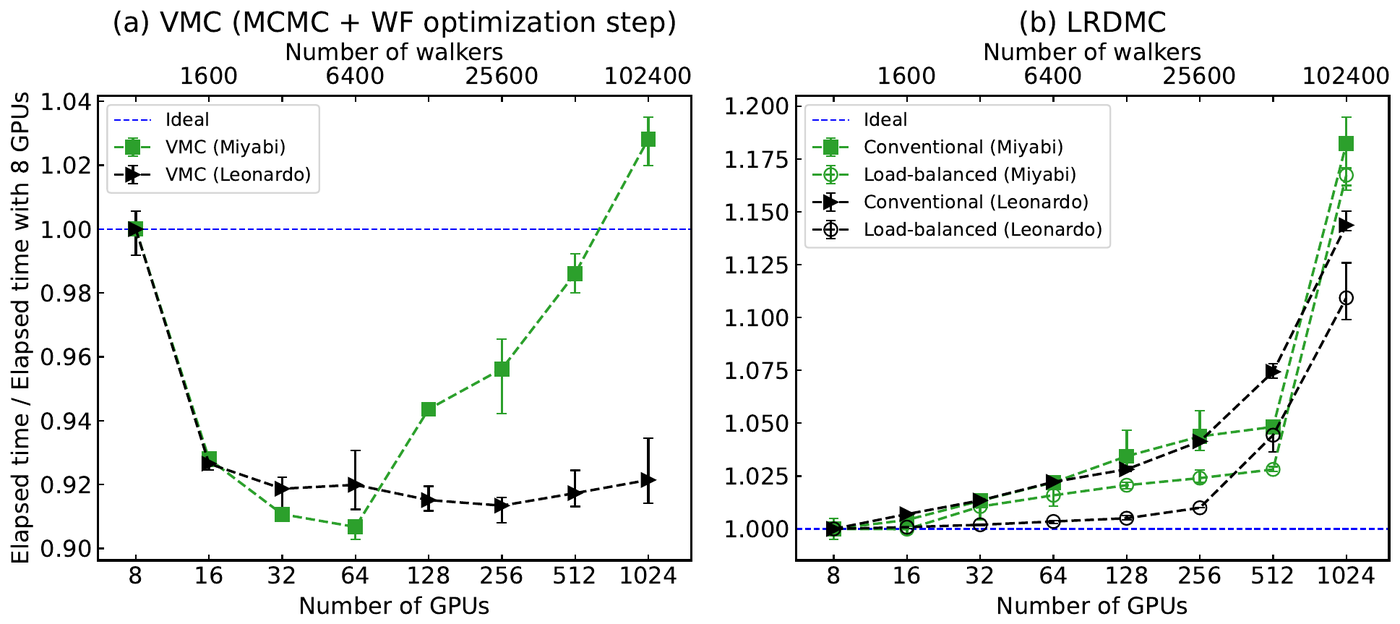
Weak-scaling benchmark of jQMC for (a) VMC and (b) LRDMC calculations, measured on Miyabi (NVIDIA H100, the University of Tokyo) and Leonardo (NVIDIA A100, CINECA) from 8 up to 1024 GPUs. jQMC retains nearly ideal weak scaling over the whole range, i.e., it sustains its parallel efficiency up to 103 GPUs. Reproduced from K. Nakano and M. Casula, arXiv:2607.13781.
Current and future research directions include:
Methodological developments in predicting material properties with QMC, particularly force calculations and their implementation into jQMC and TurboRVB.
Establishing a database of localized (Gaussian) basis sets optimized for periodic boundary conditions in QMC calculations.
Applying materials informatics approaches through screening calculations.
Constructing and applying (machine-learning) molecular dynamics force fields based on QMC-calculated data.
Some examples of applications:
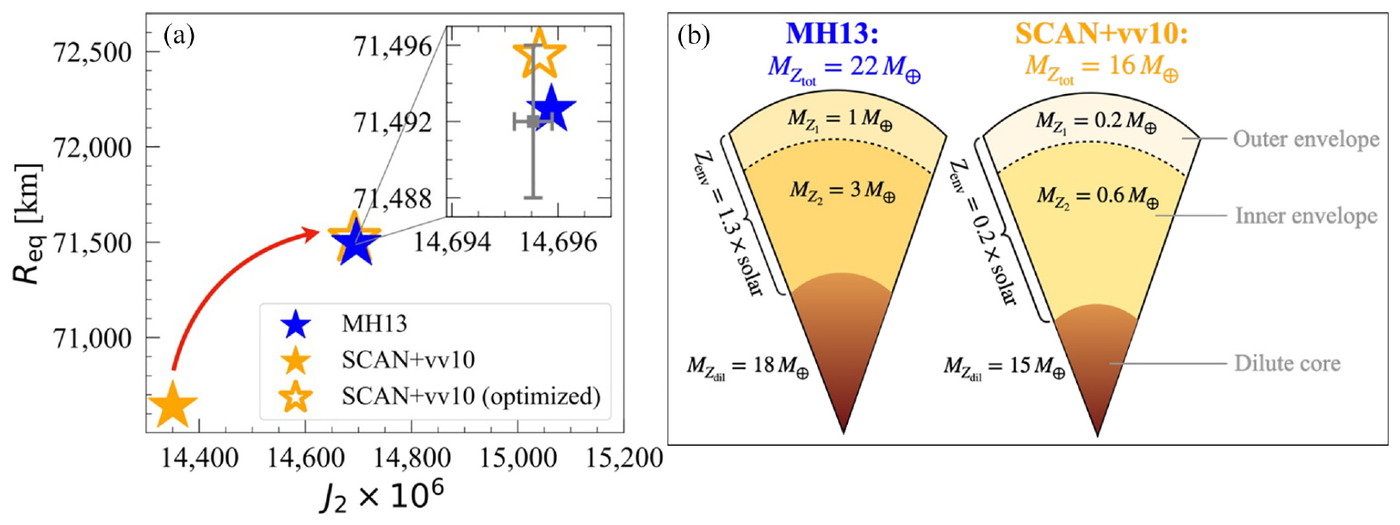
Impact of an ab initio equation of state (EoS) of hydrogen, benchmarked against our QMC calculations, on the interior structure of Jupiter. (a) Equatorial radius and gravitational moment J2 of Jupiter interior models, and (b) the resulting internal structures. A denser hydrogen EoS implies a significantly smaller heavy-element mass, which challenges the current interior models of the planet. Reproduced from C. Cozza, K. Nakano, S. Howard, H. Xie, R. Helled, and G. Mazzola, Phys. Rev. Research 8, 013089 (2026), licensed under CC BY 4.0.
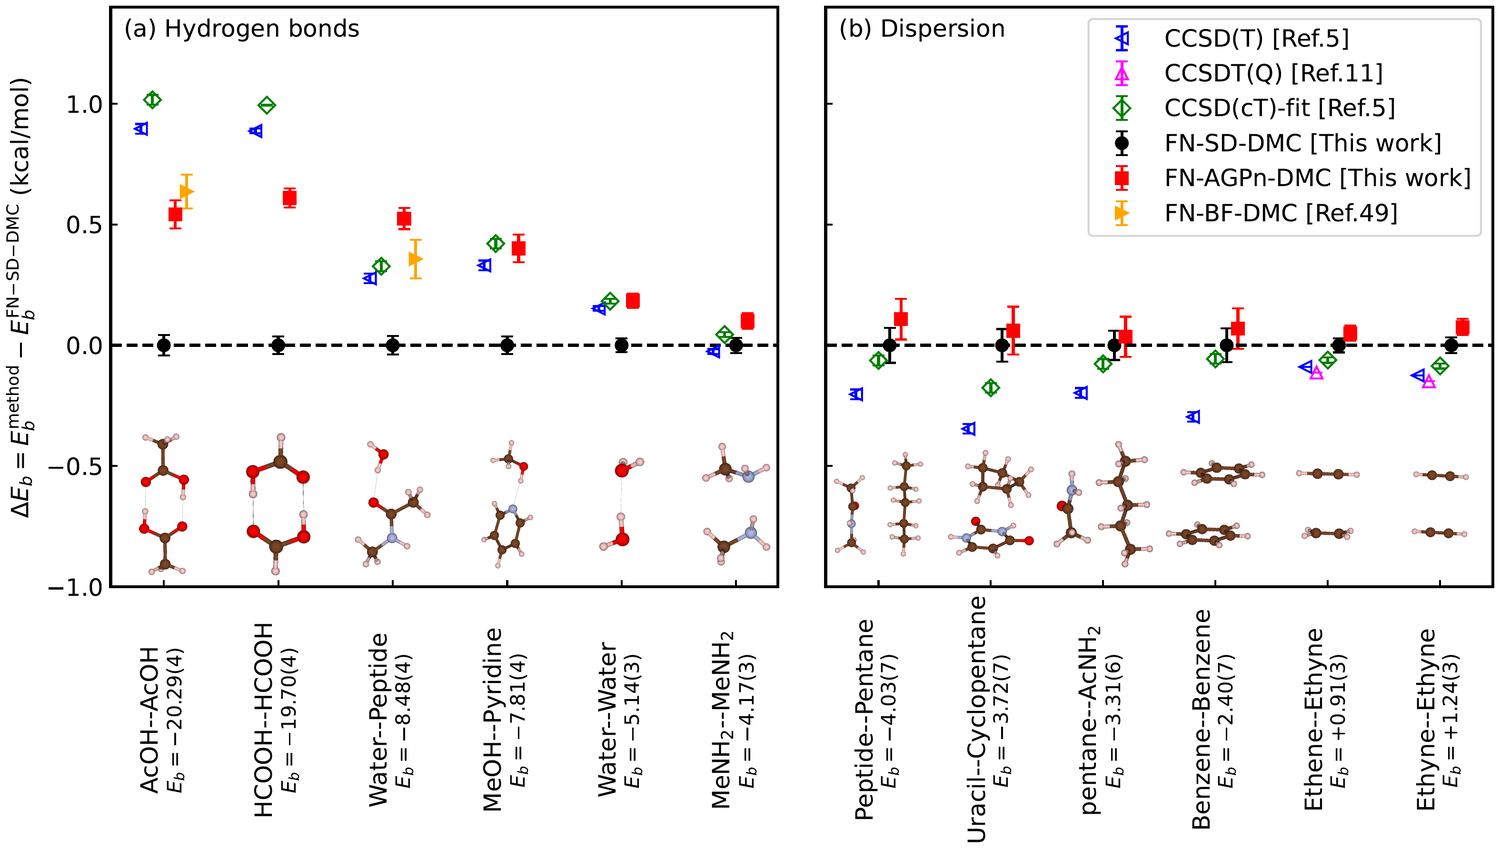
Which is more reliable for non-covalent interactions, CCSD(T) or diffusion Monte Carlo? Binding energies of (a) hydrogen-bonded and (b) dispersion-dominated dimers, plotted as the difference from our FN-SD-DMC values. Improving the nodal surface with the antisymmetrized geminal power ansatz (FN-AGPn-DMC) removes most of the discrepancy with CCSD(T) for hydrogen bonds, whereas for dispersion-dominated bonds the discrepancy survives and remains an open question. Reproduced from K. Nakano, B. X. Shi, D. Alfe, and A. Zen, J. Chem. Phys. 165, 124102 (2026), licensed under CC BY 4.0.